Rose is a globally significant ornamental crop and an emerging genomics system for woody ornamental plants. In recent decades, the rapid evolution of high-throughput sequencing and robust technical platforms has yielded high-quality reference genomes and been propelling our understanding of rose biology to unprecedented depths. This review systematically synthesizes the current landscape of rose genomics, recent development of technical and resource platforms, and the molecular mechanisms underlying pivotal agronomic traits such as floral development, scent biosynthesis, petal lifespan, and stress resilience. Despite these strides, high genome heterozygosity, polyploidy, and recalcitrance to genetic transformation remain significant barriers. We discuss how integrating cutting-edge technologies, such as pangenomics, single-cell transcriptomics, AI-assisted genomic selection, and precise CRISPR-based editing, can bridge the gap between fundamental research and practical applications. Collectively, this review provides a strategic roadmap for accelerating the development of next-generation rose cultivars through trait-based biobreeding.
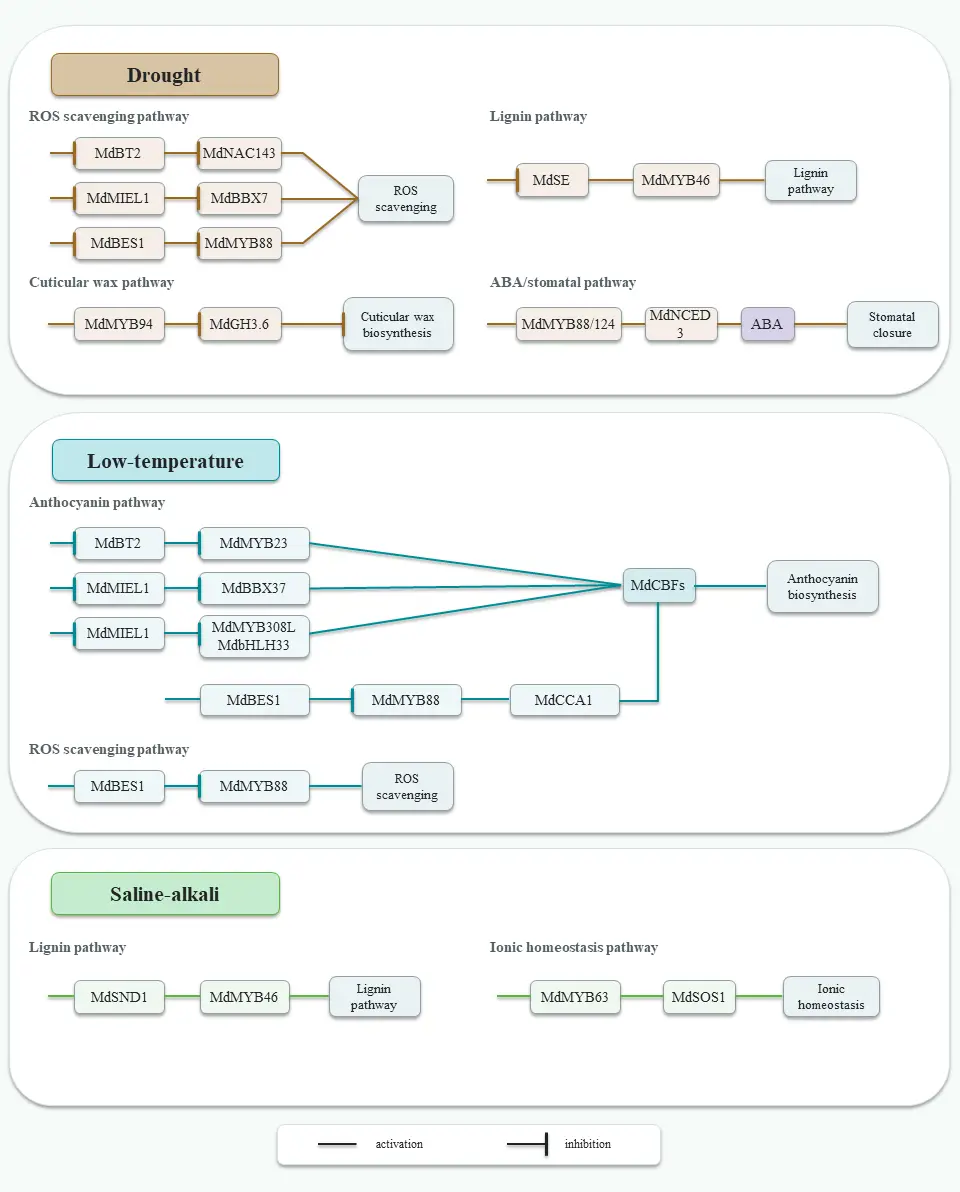
As the leading global producer of apples, China’s apple industry faces substantial challenges posed by abiotic stresses. Consequently, it is imperative to carry out an in-depth synthesis and refinement of the unique physiological and molecular mechanisms underlying apple stress resistance, as well as to comprehensively and precisely uncover their response patterns under diverse abiotic stress conditions. Such endeavors are crucial for fostering the sustainable development of the apple industry. Presently, research on abiotic stresses in Chinese apples is intricately linked to industrial issues prevalent in major apple-producing regions, with a primary emphasis on improving drought, cold, and salt-alkali tolerance. This review synthesizes studies on Chinese apples, spanning tree growth, physiological biochemistry, and molecular regulation. Key questions and future directions are outlined to inform research on stress resistance and precision breeding strategies.
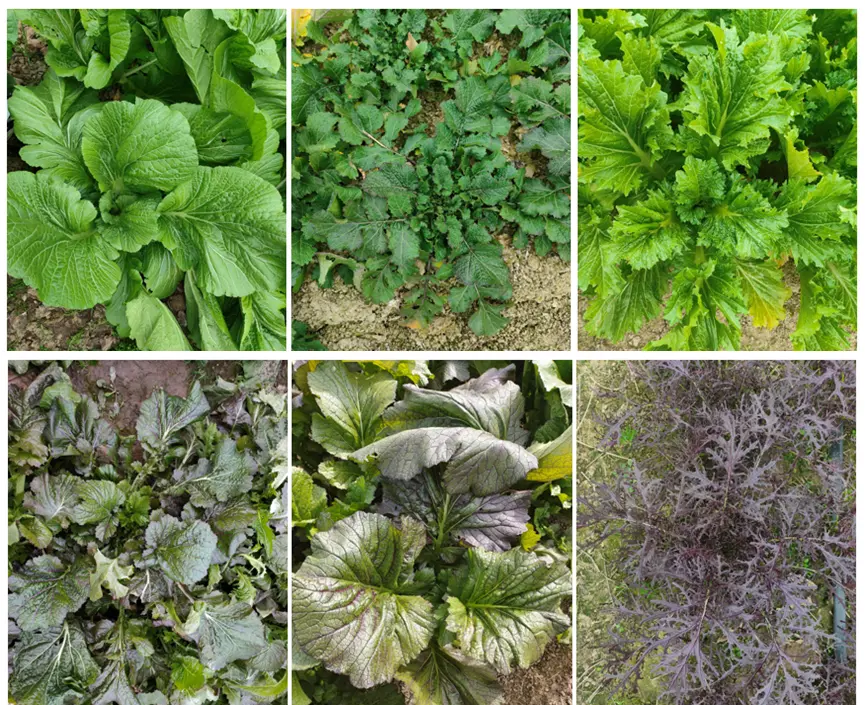
Leaf mustard (Brassica juncea) is an important leafy vegetable highly valued for its diverse flavor and nutrient compounds, particularly glucosinolates. However, different varieties of leaf mustard exhibit substantial phenotypic variation and varying glucosinolate content. In this study, we systematically assessed the phenotypic variability and glucosinolate content among 86 genotypes of leaf mustard. The Shannon-Wiener index for qualitative traits ranged from 0.10 (leaf surface gloss) to 1.60 (leaf shape), while for quantitative traits, it ranged from 1.85 (petiole length) to 2.07 (leaf width). Cluster analysis grouped the accessions into three distinct clusters, with hierarchical clustering indicating that yield-related traits were the primary factors distinguishing these groups. Principal component analysis (PCA) revealed that eight components accounted for 87.44% of the total variance in yield and glucosinolate attributes. Based on comprehensive scoring, the top five genotypes (A645, A464, A512, A702, and A445), exhibiting diverse characteristics, were identified as promising candidates for breeding programs. Moreover, our results suggest that leaf mustard leaves with deeper or more numerous lobes contain higher concentrations of glucoraphanin. Collectively, these findings provide valuable genetic resources to advance leaf mustard breeding.
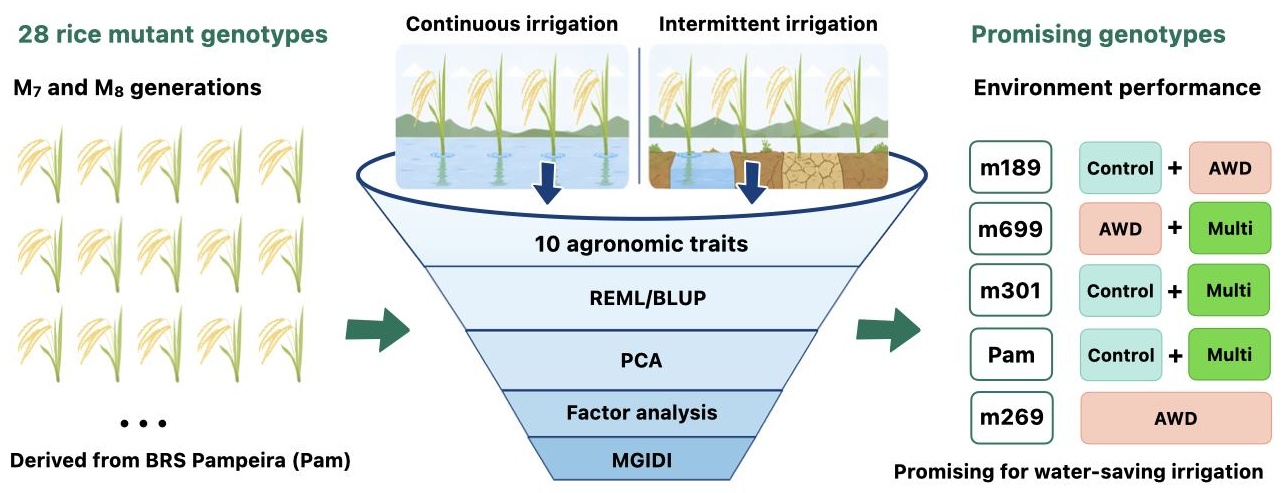
Genotype × environment (G×E) interactions complicate the identification of stable, high-performing genotypes in plant breeding, particularly under increasing climate variability and water scarcity. To support the development of water-efficient rice cultivars, we evaluated 28 mutant lines derived from the cultivar BRS Pampeira under two irrigation regimes: continuous flooding and alternate wetting and drying (AWD). Agronomic traits were assessed using mixed models (REML/BLUP) to estimate genotypic values, while principal component analysis (PCA) and the multi-trait genotype–ideotype distance index (MGIDI) were used to support genotype selection. Significant G×E interactions were observed for yield-related traits, demonstrating differential responses to irrigation regimes. The MGIDI index identified superior genotypes according to selection scenarios, with m189 showing recurrence under both individual scenarios, m699 selected under AWD and in the multi-environment analysis, and m301 and BRS Pampeira selected under Control and multi-environment conditions. Under AWD, m269 was selected as a promising genotype due to its proximity to the ideotype and balanced multi-trait performance. These findings highlight substantial genetic variability among mutant lines and provide promising candidates for improving rice productivity and adaptation under water-limited production systems.
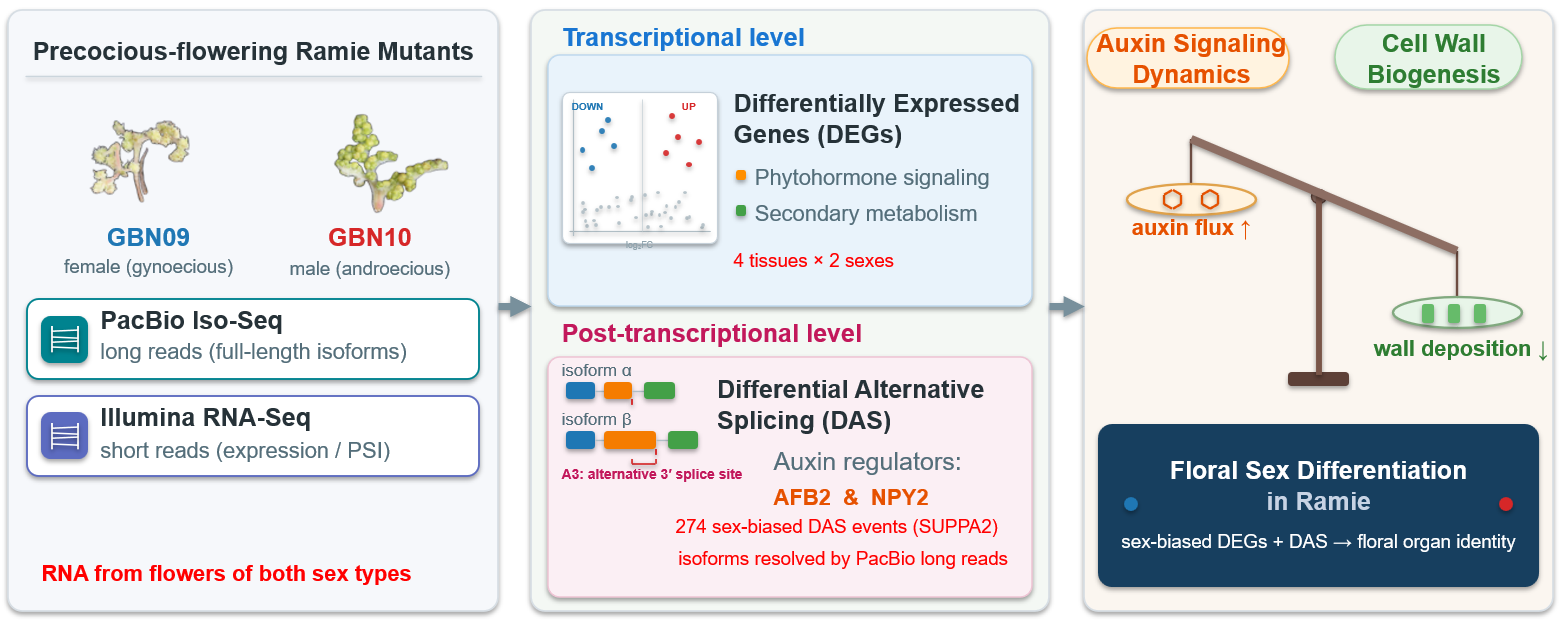
Ramie (Boehmeria nivea L.) is an economically valuable bast fiber crop, yet a high-quality full-length transcriptome reference for sex-divergent lines remains lacking. To fill this gap, we combined PacBio Iso-Seq with Illumina RNA-Seq to construct a full-length transcriptome for the gynoecious mutant GBN09 and compared transcriptomes of four tissues (leaves, bast fibers, stems, and flowers) between GBN09 and the androecious mutant GBN10. The analysis generated 63,082 high-quality isoforms and annotated 30,885 genes. Regulatory element mining revealed 565 transcription factors (predominantly IAA, AP2/ERF, and WRKY families), 16,617 long non-coding RNAs (lncRNAs), and 31,759 simple sequence repeats (SSRs). Transcriptomic comparisons identified 8414 differentially expressed genes (DEGs), with the most profound transcriptional shifts in floral and young stem tissues. Enrichment analysis linked these DEGs primarily to linoleic acid metabolism, phytohormone signaling cascades, and secondary metabolite biosynthesis, highlighting their roles in sexual differentiation. Furthermore, we detected 274 sex-biased differential alternative splicing (DAS) events, many of which mapped to cell wall biogenesis and core auxin signaling components such as AFB2 and NPY2. These results indicate that post-transcriptional modifications provide an additional regulatory layer during ramie floral sex differentiation. Validated by qRT-PCR, this dataset establishes a robust molecular resource and identifies candidate genes to elucidate sex differentiation and support molecular breeding in ramie.